Recombination Rate Is Spatially Correlated with W→S DAF Skew across the Genome
Recombination Rate Is Spatially Correlated with W→S DAF Skew across the Genome
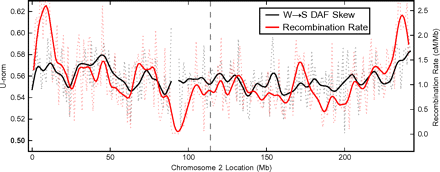
To further explore the relationship of W→S DAF skew and recombination, we correlated the W→S DAF skew in windows across the genome with the estimated recombination rate for each window for the YRI population from HapMap (The International Hapmap Consortium 2007). At a 40 kb window size, the raw Spearman correlation across the entire genome is 0.20 (P ≈ 0). When larger 1 Mb windows are considered, the correlation becomes much stronger (ρ = 0.53; P ≈ 0). Figure 5 illustrates this strong spatial correlation between bias and recombination rate across chromosome 2 (ρ= 0.43; P ≈ 0). The dramatic increase in correlation with increasing window size is consistent with models of the evolution of recombination hot spots in which there is a large amount of local variation between individuals and over time, but this variation is mainly contained within larger-scale regions with fairly constant recombination rates (Myers et al. 2005).

W→S fixation bias and recombination rate are significantlycorrelated across the genome. The W→S DAF skew (black) (asdescribed in fig. 2) and YRI population–based recombination rate (red) are significantly correlated (ρ = 0.62 smoothed and ρ = 0.43 unsmoothed; both P ≈ 0) across chromosome 2. Raw W→S DAF skew and recombination rate data in 1 Mb windows are plotted with dotted lines, and the data smoothed using a Hanning window of size 12 Mb are shown with solid lines. The dashed vertical line indicates the location of the fusion of two ancestral chromosomes on the human lineage.

没有评论:
发表评论